
Quality and Regulatory Consultancy
Helping medical device teams navigate US FDA, EU MDR and UK MDR requirements.
Hands-on regulatory and quality support for hardware, software and AI‑enabled devices, from early strategy to submission and market authorisation.
experience from UL Solutions, Intertek and Philips.
Regulators expect more
than most teams plan for.
Most regulatory delays come down to unclear expectations, incomplete technical files and evidence gaps found too late.
“We’re not sure which device class or regulatory pathway applies.”
Strategy
“We need to comply with ISO 13485, but don’t know where to start.”
Quality Management System
“How much clinical evidence is required?”
Documentation
“Our submission came back with an Additional Information request.”
Submissions
2 in 3
FDA 510(k)s get an Additional Information request in their first review.
FDA MDUFA report, FY2022–24
65%
of MDR files reach Notified Bodies missing at least half the required information.
Team-NB survey, 2025
91%
of medtech SMEs find it hard to hire qualified regulatory staff.
MedTech Europe survey, 2024
We’ve been in your shoes.
We combine quality and regulatory expertise with hands-on engineering across medical systems, electronics, software, cybersecurity and AI/ML. We work as an extension of your team, because we know what it takes to be part of one.
As
manufacturers
We’ve built devices inside a global manufacturer, so we know the pressures your team is under.
As
engineers
We’ve worked hands-on with software, electronics and AI, so nothing gets lost in translation.
As
testers
We’ve tested devices for certification, so we know what reviewers and auditors look for.
As
consultants
We’ve guided teams through UK, EU and US FDA submissions, so you know what to expect.
Engineers who speak regulatory.
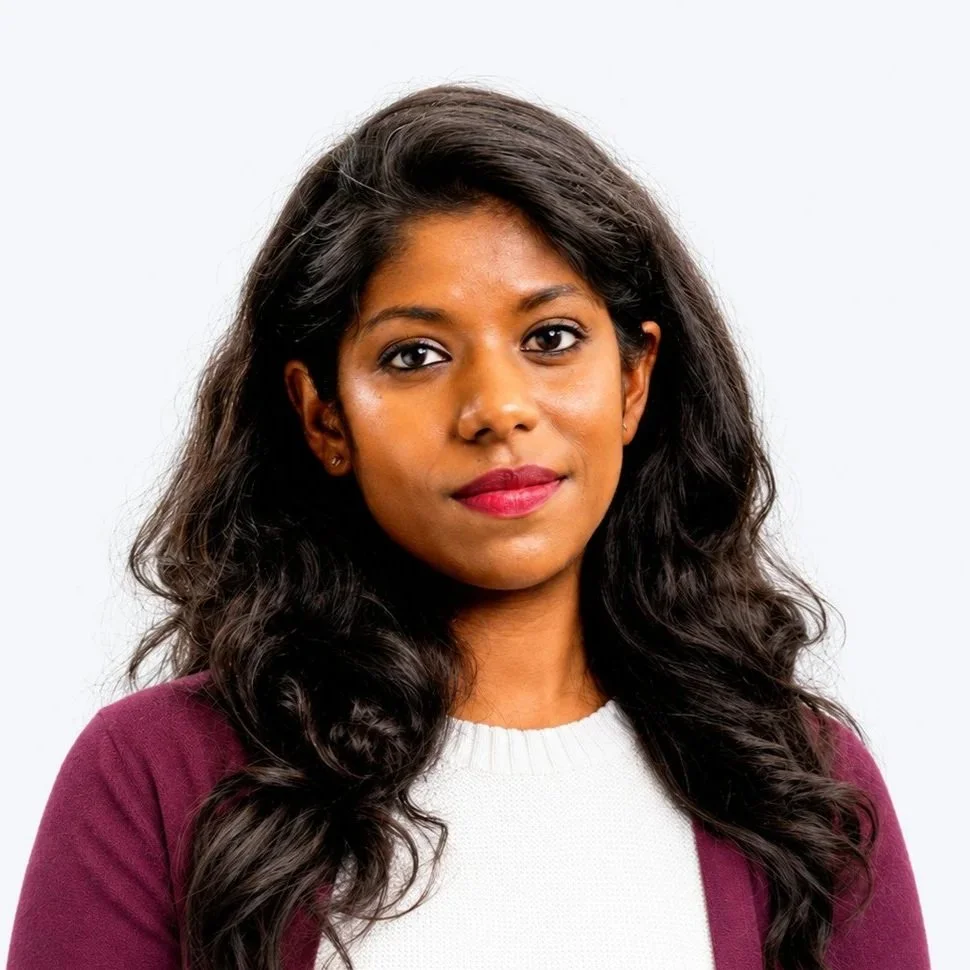
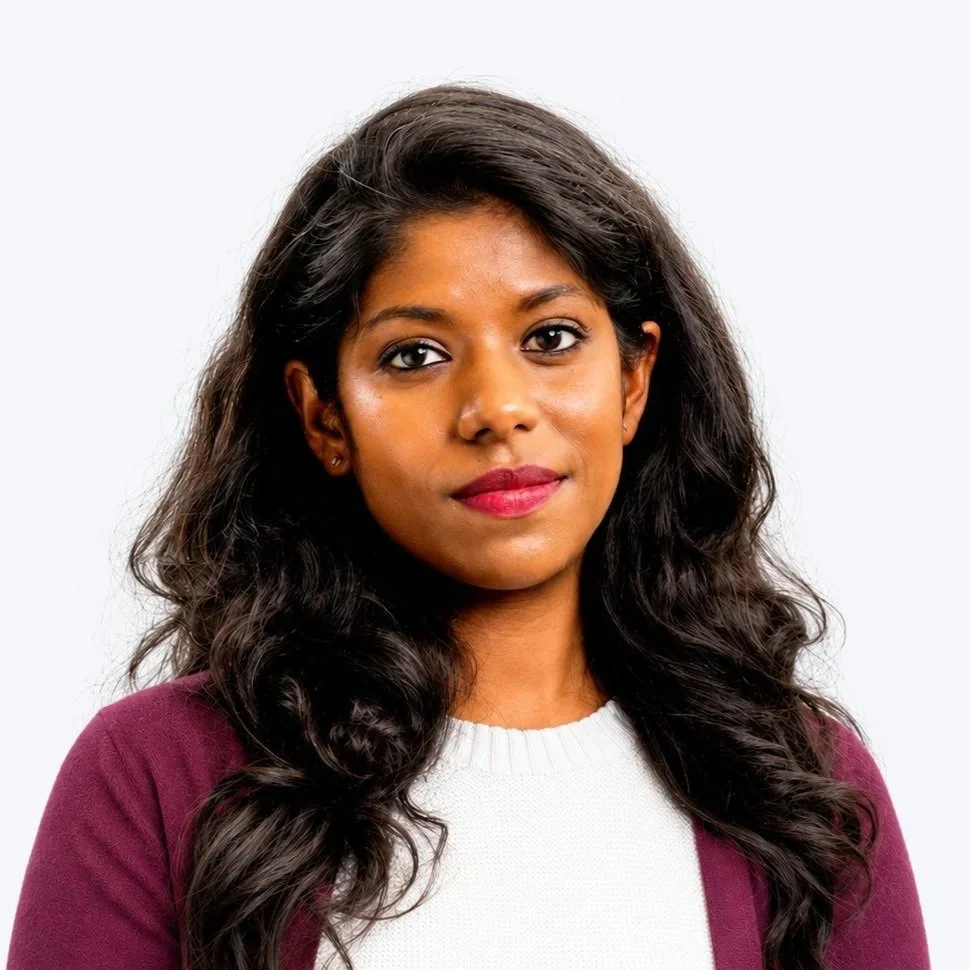
Hima Chetty
CEO and Principal ConsultantGuides every engagement from strategy to submission. Over a decade of experience in safety testing, risk management and systems engineering.
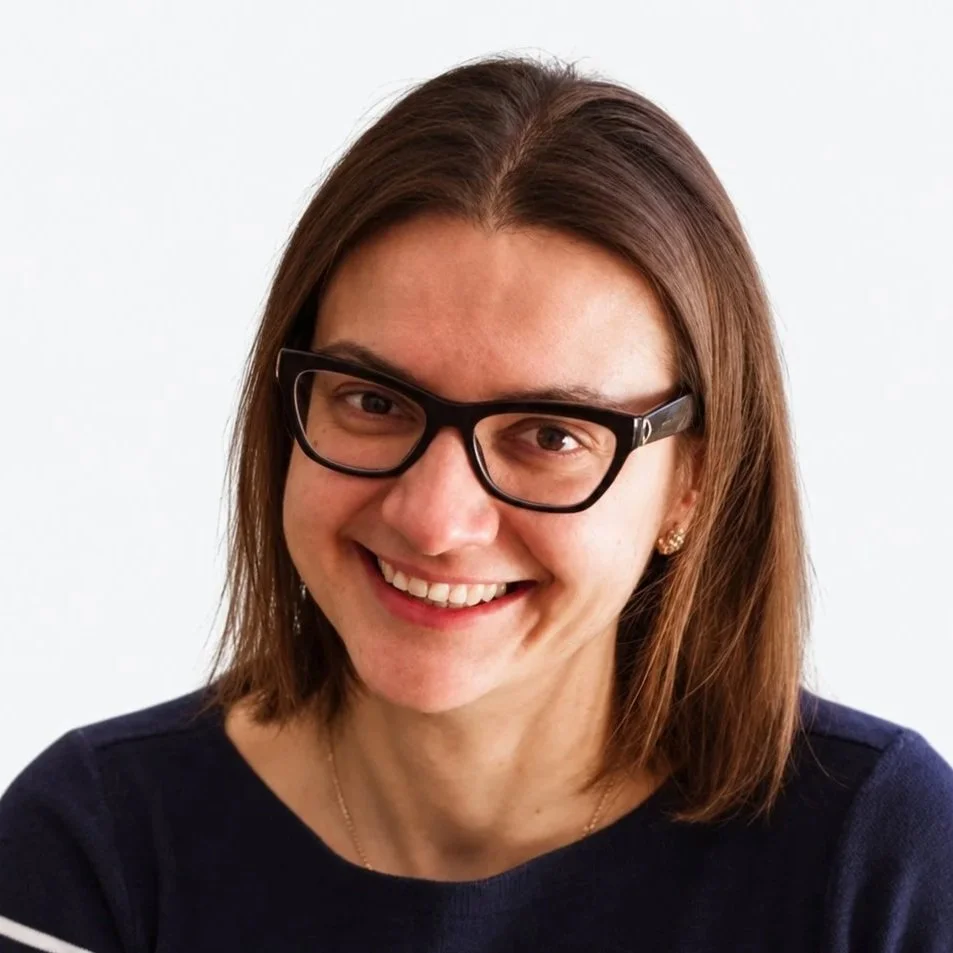
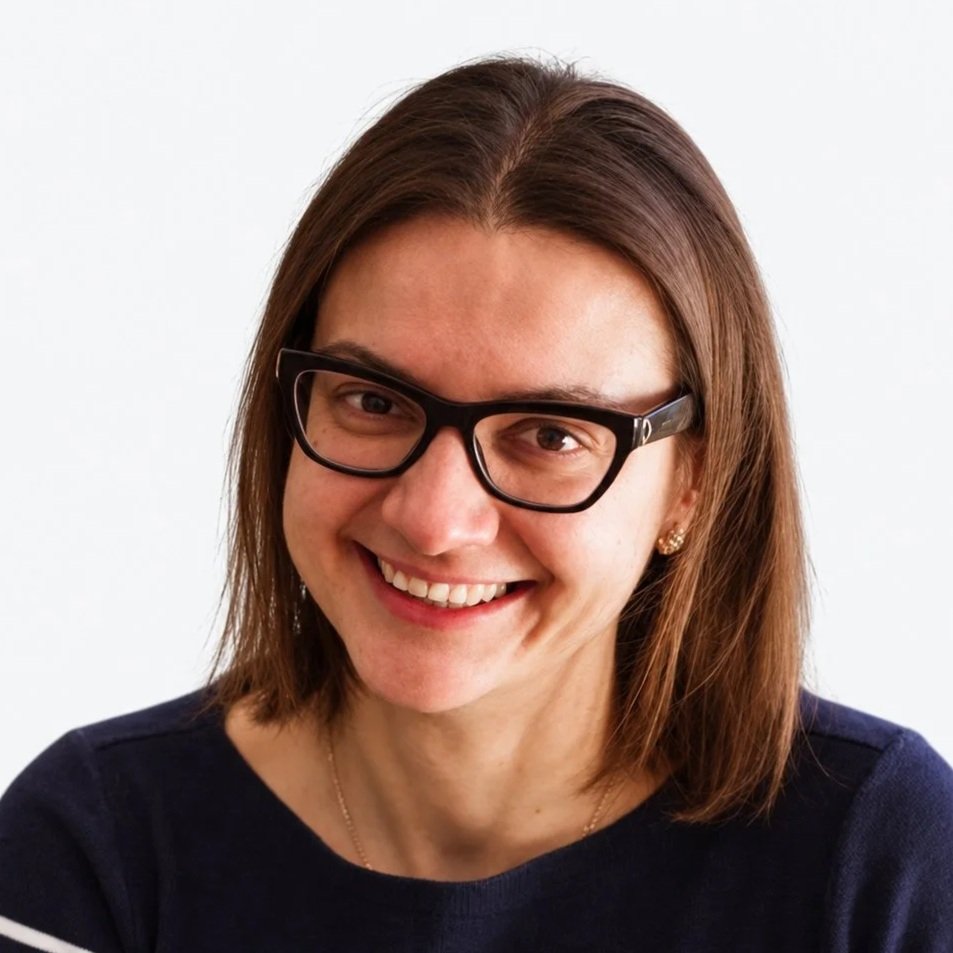
Victoria Zubova
Senior Software ConsultantLeads software, usability and risk documentation. Brings over a decade of medical device software development experience at a global manufacturer.
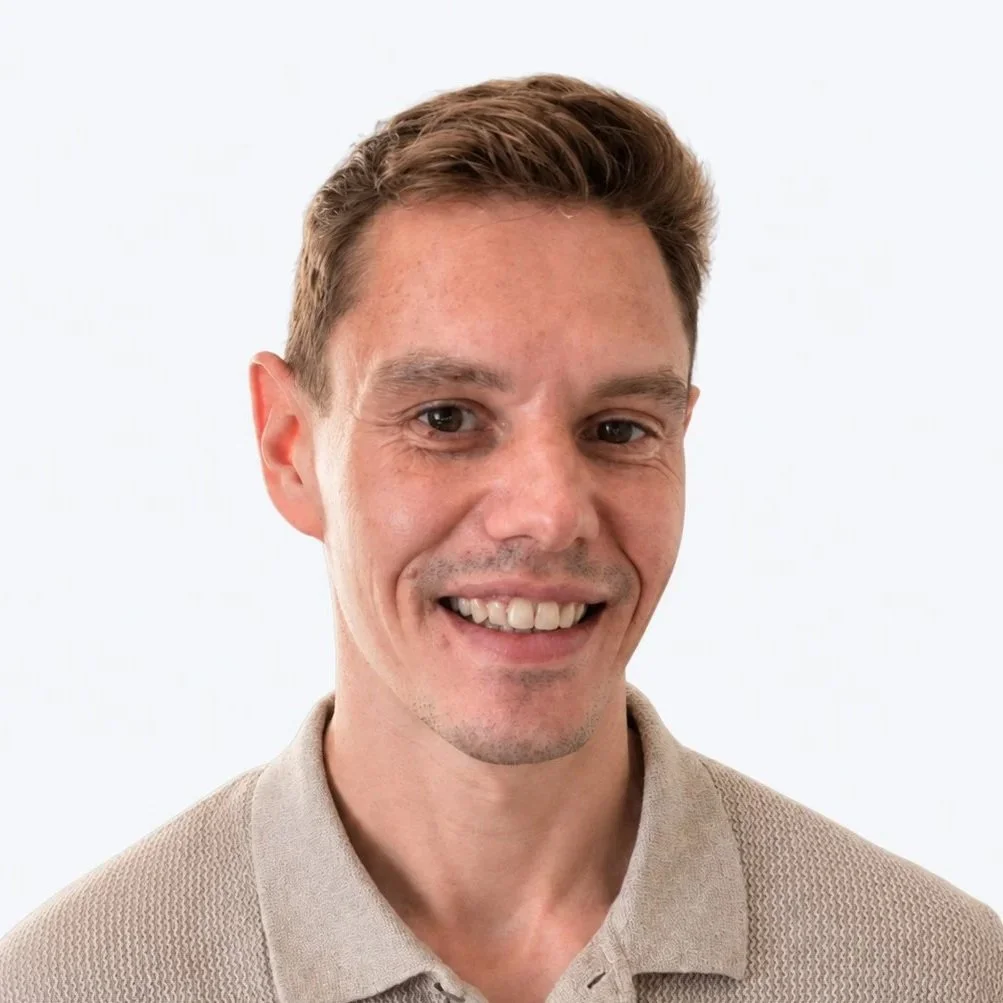
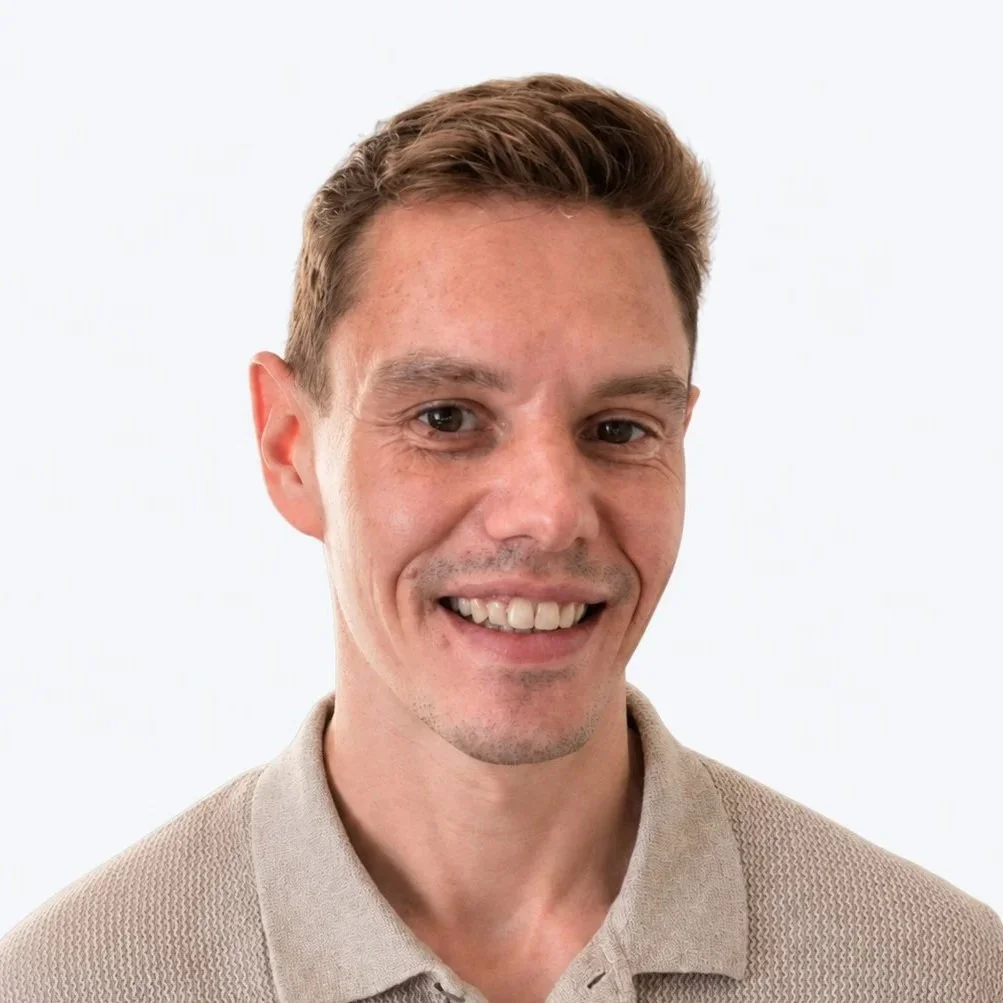
Pavel Vasilyev
Director of Operations
Keeps projects on track, with a shared tracker, regular meetings and transparent reporting. Background in delivering regulated programmes.
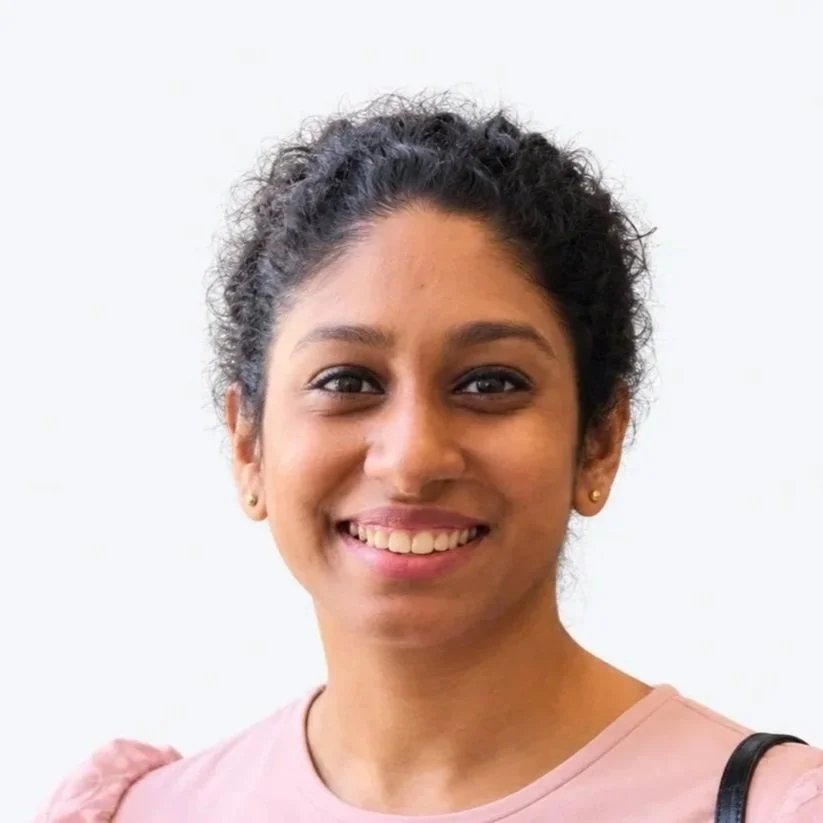
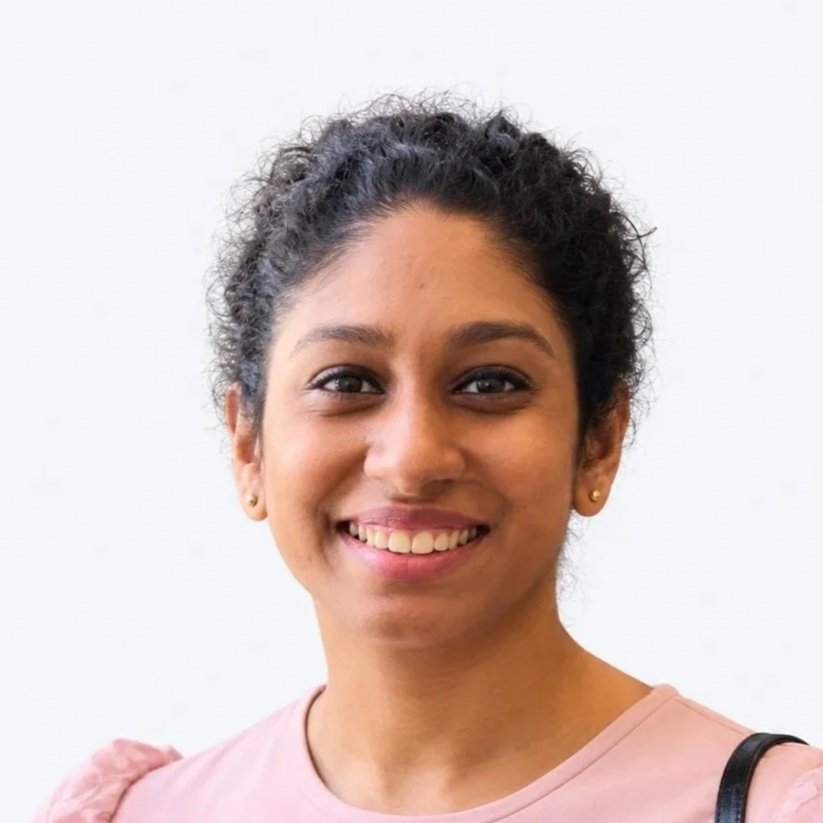
Swarna Chetty
AI/ML ConsultantHelps teams build the evidence regulators expect for AI‑enabled devices, from data management to model validation. PhD in Applied machine learning (ML).
From strategy to market authorisation.
Find the fastest route to every market
Avoid the guesswork. Find out exactly which regulations, standards and market requirements apply to your device before you commit.
- Classification and intended use
- Pathway and evidence plan per market
- Roadmap with timelines and costs
A QMS that grows with your company
Built in stages as your organisation grows. The essentials for controlled development first, then the full ISO 13485-compliant Quality Management System.
- Gap assessment and QMS design
- Procedures, design controls and training
- Internal audit and certification readiness
Technical files done your way
Regulatory-grade documentation that stands up to scrutiny. We draft for you, with you, or review yours.
- Design controls and risk management
- Software, AI/ML and cybersecurity
- Usability, labelling and post-market
Submissions built for compliance
Started on day one and built up as documentation and evidence become available, so the final package is complete and consistent.
- UK MDR registration and technical file
- EU MDR technical file and Notified Body support
- FDA 510(k), De Novo and Premarket Approval
As much support as you need.
Every team starts from a different place. Choose how involved we are, and change it document by document as your project moves on.
You lead
Review
For teams who have the expertise in-house. You draft, we review against the requirements and show you exactly what to fix before an auditor does.
We work together
Collaborate
For teams who want to build capability as they go. We draft side by side in working sessions, so your people learn the reasoning behind each decision.
We lead
Done for you
For teams who need to move fast without hiring. We lead the work from your inputs, keep you informed at every step, and you review and sign off.
We help medical device teams bring safe, effective technology to patients sooner. We bring engineering depth and regulatory rigour to every project, and we work as part of your team, not from the sidelines.
Questions we often hear.
Can’t find what you’re looking for? We’re here to help and answer any questions you may have.
-
Medical device, software as a medical device and AI-enabled device teams, from early-stage start-ups preparing their first submission to established manufacturers entering a new market.
-
The UK (UK MDR and MHRA registration), the EU (EU MDR and Notified Body certification) and the United States (FDA 510(k), De Novo and Premarket Approval). We often plan all three together, so evidence is generated once and reused.
-
Yes, that’s where many of our projects begin. We review what you already have, list the gaps in order of risk, and pick up from there without redoing work that's already sound.
-
Yes. With our Done for you option we lead the work, and you provide the device knowledge and test results only your team has. As the manufacturer, you stay responsible for the final sign-off, and we make that step simple.
-
It depends on the device class, the market and how much evidence already exists. A UK Class I registration can be a matter of weeks; an EU MDR Class IIa certification or an FDA 510(k) usually takes months, much of it in review. Your roadmap includes a realistic timeline from the start.
-
Send us a short note about your device and target markets. We'll arrange a call, then send a proposal with scope, estimates and timeline.
Let’s plan your
route to market.
Tell us about your device and where you want to sell it. We’ll come back with a clear next step, whether that's a quick answer or a full roadmap.